TLDR
For medical device companies preparing for clinical production or commercialization, ethylene oxide (EO) sterilization can introduce substantial validation, testing, scheduling, and documentation requirements. However, not every device necessarily requires an entirely new sterilization cycle. Under appropriate circumstances, a new or modified product may be eligible for adoption into an existing validated EO process.
ISO 11135 provides requirements for EO sterilization process development, validation, and routine control, while AAMI TIR28 addresses product adoption and process equivalence. Eligibility depends on documented evaluations of product design, materials, packaging, sterilization resistance, and load configuration. For OEMs, assessing these factors early can help determine whether an existing cycle offers a practical path forward or whether additional studies or full validation will be necessary.
Sterilization Planning Begins Before Manufacturing Is Complete
For medical device OEMs approaching design verification, clinical manufacturing, or commercial launch, sterilization can become one of the more consequential elements of production planning. The device may already function as intended, the manufacturing process may be approaching qualification, and packaging development may be underway. Yet an unresolved sterilization strategy can affect when finished units become available for their intended use.
Ethylene oxide sterilization is widely used for medical devices because it can accommodate many materials and configurations that may not tolerate other sterilization methods. Catheters, tubing assemblies, procedure kits, and devices incorporating temperature-sensitive components are among the product categories commonly processed using EO. Its versatility, however, does not eliminate the need to establish and document an appropriate sterilization process for the specific device and its production configuration. STERIS AST
One question that frequently arises during commercialization planning is whether the device can be incorporated into an existing validated EO sterilization process rather than requiring a newly developed cycle. The potential benefits are meaningful: a suitable existing process may reduce redundant development work, simplify certain qualification activities, and provide a more established operational framework. Those benefits are only achievable when the manufacturer can demonstrate that the new product is appropriately covered by the validated process.
For companies transitioning from engineering prototypes into regulated manufacturing, the decision should be evaluated alongside material selection, device assembly, packaging development, quality documentation, and expected production volumes. Waiting until the final stages of commercialization to address sterilization compatibility can introduce additional testing and changes at a point when the program has limited flexibility.
What Does an Existing Validated EO Cycle Actually Cover?
A validated EO sterilization cycle is a documented process established to achieve defined sterilization requirements under specified operating conditions. ISO 11135:2014, including its 2018 amendment, establishes requirements for the development, validation, and routine control of EO sterilization processes for medical devices. As of October 2026, ISO 11135:2014 remains the published international standard, although a revision is under development. ISO
EO sterilization involves controlled process variables, including temperature, humidity, sterilant concentration, exposure time, and applicable load characteristics. These variables influence whether the sterilization process can effectively address the microbiological challenge presented by the product. The validation establishes an evidence-based operating framework, but that framework is not automatically applicable to every medical device placed into the chamber.
A previously validated process may accommodate multiple devices or product configurations when the technical justification supports their inclusion. Such products are commonly organized into sterilization families based on characteristics that influence their response to the process. Products within a family may differ in dimensions or configuration, but their inclusion must be supported by appropriate evaluation of the validated process and its established challenge conditions.
This distinction is important for procurement and operations teams. Access to an existing validated cycle is not equivalent to having an automatically approved sterilization method for a new device. A manufacturer must determine whether the candidate product falls within the demonstrated capabilities and limitations of that process.
Understanding EO Sterilization Product Adoption
Product adoption is the process of evaluating whether a new or modified medical device can be incorporated into an existing validated sterilization process. AAMI TIR28:2016, reaffirmed in 2024, addresses product adoption and process equivalence for EO sterilization. It provides a framework for assessing relevant product changes, differences between products, and the evidence needed to support an adoption decision. AAMI
The evaluation generally begins by comparing the candidate device with the existing validated product family or applicable process challenge device. Relevant differences may include device geometry, material composition, packaging characteristics, density, and the physical configuration of the sterilization load. The assessment considers whether those differences introduce a challenge beyond what the existing process has already demonstrated it can accommodate.
For example, an OEM developing a revised catheter configuration may initially assume that the previous product’s sterilization cycle remains suitable. If the new configuration incorporates additional lumens, different polymers, a modified sterile barrier system, or a substantially different load arrangement, those changes require assessment. Even where the finished device appears similar to the original product, changes in construction can influence sterilant penetration, process response, and residual retention.
The outcome may be a documented justification supporting adoption, a determination that additional studies are needed, or a conclusion that the existing process does not adequately cover the proposed product. The appropriate path depends on the technical evidence rather than the similarity of product names or the commercial convenience of using the same sterilization provider.
Four Factors That Influence EO Product Adoption
1. Device Design and Geometry
The physical construction of a medical device influences its potential resistance to sterilization. Internal lumens, enclosed spaces, overlapping components, restricted openings, and complex assemblies can affect how sterilant reaches areas that require sterilization. These characteristics become particularly relevant for products such as catheters, fluid delivery systems, and devices incorporating multiple interconnected components.
A design modification that appears minor from an engineering perspective may create a meaningful change in sterilization challenge. Increasing tubing length, reducing lumen diameter, or introducing a new assembled interface may require further evaluation. Conversely, changes that do not introduce a greater sterilization challenge may be easier to justify within an existing validated product family, provided the assessment is appropriately documented.
2. Material Composition
Medical devices frequently combine polymers, metals, adhesives, coatings, and other specialized materials. These materials can respond differently to EO exposure and subsequent aeration. Material composition may influence sterilant absorption, residual retention, and device performance following processing.
For OEMs, this means sterilization compatibility cannot be evaluated solely on the basis of whether EO will damage a material. The manufacturer must also consider whether the finished device retains acceptable physical, chemical, and functional characteristics after processing and whether residual sterilant levels can be appropriately controlled.
3. Sterile Barrier Packaging
The sterile barrier system affects how sterilant reaches the device and how sterility is maintained after processing. Packaging materials, pouch construction, tray geometry, protective inserts, and overall package dimensions may influence the sterilization process.
A new device packaged in a substantially different configuration from an existing product may require additional evaluation even when the underlying device materials are similar. EO compatibility, package integrity, sealing-process controls, and post-sterilization package performance should therefore be considered together rather than evaluated as unrelated manufacturing activities.
4. Load Configuration and Density
Sterilization validation also depends on the physical configuration of products inside the chamber. Carton size, pallet arrangement, load density, and the distribution of materials can influence temperature and humidity conditions and sterilant penetration.
A process validated using one loading configuration does not necessarily cover a substantially different arrangement. A new device may therefore require evaluation of both the product itself and the intended commercial load, particularly when production volumes or packaging formats differ from those used during the original process qualification.
These factors demonstrate why product adoption is a technical assessment rather than an administrative shortcut. STERIS identifies product design, materials, sterile barrier configuration, and loading characteristics as central considerations when evaluating whether a product can enter an existing validated EO process.
EO Residuals Are a Separate Consideration
Demonstrating that a sterilization process achieves its intended microbiological performance is not the only requirement for EO-sterilized medical devices. Residual sterilant and relevant reaction products must also be assessed where applicable to the device and its intended use.
This subject became particularly relevant in 2026. ISO published ISO 10993-7:2026 in April 2026, replacing ISO 10993-7:2008. The updated standard addresses allowable limits for residual ethylene oxide and ethylene chlorohydrin, methods for measuring those substances, and approaches for determining conformity. Manufacturers should evaluate the current edition and applicable regulatory recognition requirements when developing their biological evaluation and residual-testing strategies. ISO
Material composition, exposure conditions, device geometry, and aeration can influence residual behavior. A product considered suitable for adoption from a sterilization-resistance standpoint may still require evaluation of whether acceptable residual levels can be achieved under the proposed processing conditions.
This distinction becomes especially important for devices incorporating absorbent materials, multiple polymer types, adhesives, or configurations that may retain sterilant differently from existing products. Planning for residual evaluation early helps manufacturers avoid treating sterilization processing and final product release as the same milestone.
Where a manufacturer intends to rely on a voluntary consensus standard in an FDA premarket submission, the edition and scope of FDA recognition should also be checked. Publication of a new ISO edition does not, by itself, establish that the FDA recognizes that edition for a declaration of conformity.
Packaging Validation and Sterilization Must Work Together
A medical device may be compatible with EO sterilization while its packaging configuration still requires additional development or testing. This is why packaging and sterilization strategies should be coordinated before formal validation activities begin.
ISO 11607-1 addresses requirements for materials, sterile barrier systems, and packaging systems intended to maintain sterility until the point of use. ISO 11607-2 addresses validation requirements for packaging forming, sealing, and assembly processes. Together, these standards establish important expectations for terminally sterilized medical device packaging, separate from the validation requirements governing the sterilization process itself. ISO
An OEM considering an existing EO cycle should therefore evaluate whether the proposed packaging is compatible with that process and whether the packaging system remains suitable following sterilization, handling, distribution, and storage. Relevant activities may include package integrity testing, seal-strength evaluation, aging studies, distribution simulation, and packaging process qualification.
Changes made to accommodate sterilization may also affect packaging validation. For example, modifying protective packaging to improve gas accessibility could change the way a device is supported during shipment or alter sealing requirements. Addressing these interactions early creates a better opportunity to resolve technical issues without repeating completed work.
Pathway MedTech supports sterile barrier development, ISO 11607 packaging validation, cleanroom packaging, aging and distribution-related testing, and sterilization coordination. These capabilities allow packaging requirements to be considered alongside device manufacturing and the proposed sterilization approach.
When Is Additional EO Validation Necessary?
An adoption evaluation does not always lead to the same conclusion. In practical terms, an OEM may encounter three general outcomes.
The first is documented adoption without additional sterilization-cycle qualification. This may be appropriate when the technical assessment establishes that the candidate product is adequately represented by the existing validated process and the applicable evidence supports that conclusion. Other product-specific requirements, including residual evaluation, packaging qualification, and functional testing, must still be addressed as necessary.
The second is adoption supported by additional studies. If the available evidence cannot fully establish the proposed product’s suitability, targeted testing may help determine whether adoption is justified. The specific studies depend on the unresolved technical questions and the relevant sterilization requirements.
The third is a more extensive qualification or a newly validated process. Where the candidate product represents a greater challenge or cannot be appropriately accommodated within the existing process, additional qualification or development may be required. This can introduce further engineering, testing, documentation, and scheduling activities.
None of these outcomes should be assumed before the product has been evaluated. An existing EO cycle can offer significant operational advantages, but an unsupported adoption decision may create greater risks than the qualification work it was intended to avoid.
How EO Sterilization Affects Manufacturing Costs and Timelines
For an early-stage medical device company, sterilization costs extend beyond the price of processing an individual batch. The complete program may involve technical assessment, protocol development, microbiological testing, residual evaluation, packaging activities, qualification runs, documentation, transportation, and recurring production controls.
These costs can be particularly consequential for companies planning limited design verification or clinical builds. A program manufacturing a relatively small number of units may still require substantial evidence supporting the sterilization process. As a result, the relationship between nonrecurring validation expenses and ongoing processing costs should be understood before selecting a supplier or establishing a manufacturing budget.
A validated process that can accommodate a new product may offer opportunities to reduce certain development or qualification activities. However, those potential efficiencies should not be translated into guaranteed savings before the adoption assessment is complete. The cost of necessary comparative studies, product-specific testing, or process modifications may still be significant.
Production planning also matters. If clinical manufacturing, packaging qualification, and sterilization assessment occur independently, the program may encounter avoidable scheduling conflicts. Coordinating the work can help identify which decisions must be finalized before samples are manufactured, packaged, or submitted for processing.
For commercialization-stage OEMs, the relevant question is not simply which provider offers the lowest sterilization price. It is which manufacturing and sterilization arrangement provides a technically defensible, repeatable, and commercially practical process for the intended product.
Quality Responsibilities Under the FDA’s QMSR
The FDA’s Quality Management System Regulation became effective on February 2, 2026, incorporating ISO 13485:2016 by reference into the revised 21 CFR Part 820 framework. The regulation applies to manufacturers within its scope and reinforces the importance of appropriate quality management processes, documentation, supplier controls, and risk-based decision-making. U.S. Food and Drug Administration
For an OEM outsourcing manufacturing and sterilization, the responsibilities of each organization should be clearly established. The device manufacturer, contract manufacturer, contract sterilizer, and testing laboratories may each perform activities contributing to the final product, but responsibility for obtaining and maintaining appropriate evidence must be understood.
A quality agreement can help establish expectations for sterilization process changes, approved product configurations, supplier responsibilities, records, nonconformances, deviations, and product release activities. The appropriate division of responsibilities depends on the specific business arrangement and applicable regulatory requirements.
Manufacturers should also consider how sterilization-related decisions are reflected in product risk management, design documentation, manufacturing records, and change-control processes. Changes to materials, packaging, product geometry, or loading configurations may trigger reevaluation of a previously established adoption decision.
Evaluating Product Adoption Before Requesting a Manufacturing Quote
An OEM does not need to have every sterilization question resolved before approaching a contract manufacturer. However, the quality of the initial technical information can substantially affect how efficiently the program is evaluated.
A useful starting point is a clear description of the device, including its intended use, current design configuration, materials, relevant drawings, and anticipated packaging system. The manufacturer should also understand whether the device is still undergoing engineering development, approaching design verification, or preparing for recurring commercial manufacturing.
Information about expected production volume and packaging configuration is especially valuable. The number of devices per package, carton dimensions, anticipated lot quantities, and proposed pallet configuration may influence whether a product can be considered within an existing sterilization process.
OEMs should also identify any available sterilization history, material compatibility information, bioburden data, biological evaluation work, and prior performance testing. Not every program will have this information at the initial quotation stage, but identifying what exists and what remains unresolved helps distinguish preliminary assessment from formal validation activities.
The resulting discussion should clarify which activities are included in the manufacturing proposal, which require an external sterilization specialist or laboratory, and which assumptions could materially affect pricing or scheduling. This is particularly important when comparing proposals that may appear similar at the unit-cost level but contain different responsibilities for validation and documentation.
How Pathway MedTech Supports Sterilization Readiness
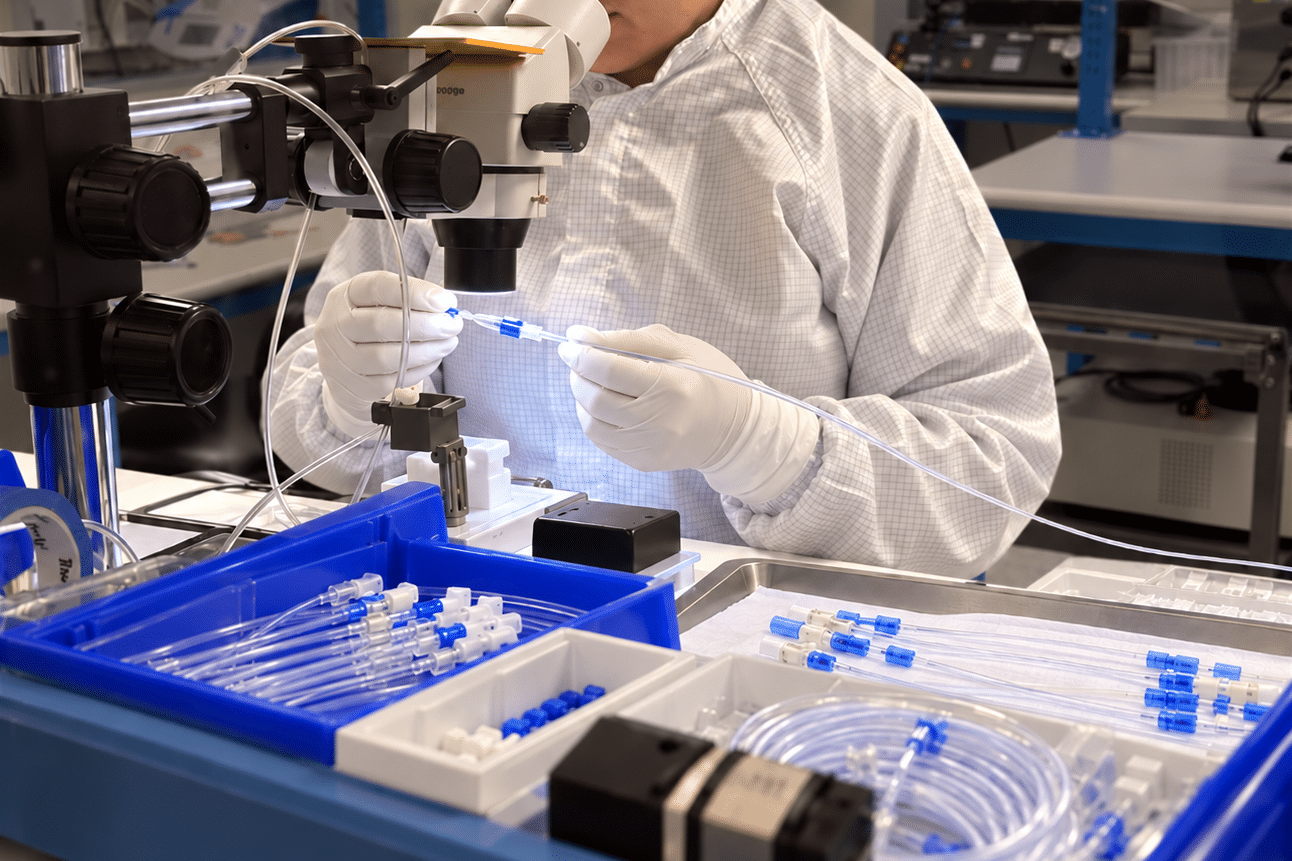
Pathway MedTech approaches manufacturing as a connected process involving engineering, quality and regulatory requirements, controlled assembly, packaging, and commercialization planning. For medical device companies transitioning from prototypes into verification or early commercial production, this creates an opportunity to evaluate sterilization requirements while the manufacturing process is still being established.
Pathway’s published capabilities include cleanroom assembly and packaging, ISO 11607 packaging validation, sterilization coordination, and access to a validated two-pallet EO family cycle through STERIS in Temecula. The availability of that established process can provide a useful starting point for evaluating eligible customer products, subject to appropriate product adoption assessment and any necessary additional testing. Pathway can also coordinate alternative sterilization approaches where required by the device and program. Pathway | New Product Development
The value of this approach is not simply access to a sterilization process. It is the ability to evaluate how sterilization requirements interact with component selection, assembly methods, cleanroom manufacturing, packaging configuration, documentation, and production scheduling.
For OEMs with limited internal manufacturing infrastructure, bringing these considerations into the project earlier can reduce the number of disconnected decisions that must be resolved immediately before regulatory submission or commercialization.
Product Adoption Should Be a Manufacturing Decision Made Early
Using an existing validated EO sterilization cycle may provide an efficient path for certain medical devices, but eligibility must be established through appropriate technical evaluation. Product design, materials, packaging, microbiological challenge, load configuration, and residual behavior all influence whether a candidate device can be supported by an existing process.
For medical device OEMs preparing for design verification, clinical production, or commercialization, the strongest approach is to evaluate sterilization alongside manufacturing and packaging development rather than treating it as a final production step. Early assessment creates the opportunity to identify whether product adoption is feasible, determine what additional evidence may be required, and incorporate realistic sterilization activities into the overall manufacturing plan.
Pathway MedTech supports this transition by connecting engineering, regulated manufacturing, packaging validation, quality and regulatory activities, and sterilization coordination. For companies evaluating an existing EO cycle or preparing to establish a new sterilization strategy, a technical review of the device and its manufacturing requirements is a practical place to begin.
Frequently Asked Questions
Can a new medical device use an existing validated EO sterilization cycle?
Yes, under appropriate circumstances. A new device may be eligible for adoption into an existing validated EO process when a documented technical assessment establishes that the process adequately covers the candidate product. The evaluation should consider product design, materials, packaging, sterilization resistance, and intended load configuration.
What is the difference between EO product adoption and EO sterilization validation?
EO sterilization validation establishes evidence supporting a defined sterilization process. Product adoption evaluates whether a new or modified product can be appropriately included within an existing validated process. An adoption assessment may support inclusion without extensive additional qualification, or it may identify studies or further validation work that must be performed.
Does EO product adoption eliminate sterilization testing?
Not necessarily. Product adoption can reduce the need for certain sterilization-cycle qualification activities when sufficient evidence already exists. However, the candidate device may still require product-specific evaluations, residual testing, packaging assessments, or other studies. The required work depends on the technical findings and applicable requirements.
What standard applies to EO sterilization product adoption?
AAMI TIR28:2016, reaffirmed in 2024, provides guidance on product adoption and process equivalence for EO sterilization. ISO 11135 establishes requirements for EO sterilization process development, validation, and routine control.
How do EO residuals affect product adoption?
Residual EO and ethylene chlorohydrin must be assessed where applicable. Material composition, device configuration, exposure conditions, and aeration may influence residual behavior. ISO 10993-7:2026 addresses allowable residual levels and related measurement and conformity assessment methods.
Can an OEM change its packaging and continue using the same EO sterilization cycle?
Possibly, but the change requires evaluation. Packaging modifications can influence sterilant penetration, load characteristics, and sterile barrier performance. The manufacturer should assess the impact on the validated sterilization process and determine whether additional packaging or sterilization studies are needed.
Can a contract manufacturer coordinate EO sterilization without operating its own chamber?
Yes. Medical device contract manufacturers can coordinate sterilization services through qualified external sterilization providers, provided responsibilities, supplier controls, process requirements, and applicable documentation are appropriately established.
When should sterilization planning begin?
Sterilization planning should begin while the device’s materials, design, and packaging requirements are being established. Early evaluation is particularly valuable before design verification or clinical manufacturing because changes to the sterilization approach can affect production configuration, testing, packaging, documentation, and project schedules.
References
- FDA: Quality Management System Regulation (QMSR). Current federal quality management requirements for medical device manufacturers, effective February 2, 2026. FDA QMSR Overview
- ISO 11135:2014. Requirements for the development, validation, and routine control of ethylene oxide sterilization processes for medical devices. ISO 11135 Standard
- AAMI TIR28:2016/(R)2024. Product adoption and process equivalence for ethylene oxide sterilization. AAMI TIR28
- ISO 10993-7:2026. Biological evaluation of medical devices, Part 7: Ethylene oxide sterilization residuals. ISO 10993-7:2026
- ISO 11607-1:2019 and ISO 11607-2:2019. Requirements for sterile barrier systems and validation of forming, sealing, and assembly processes. ISO 11607-1 | ISO 11607-2
- STERIS Applied Sterilization Technologies. Technical guidance on product adoption for ethylene oxide sterilization. STERIS Product Adoption Guidance
- FDA: Recognized Consensus Standards Database. Source for checking the FDA recognition status and applicable editions of voluntary consensus standards. FDA Standards Database