TLDR
Medical device development is governed by a combination of regulatory requirements and industry standards designed to help ensure devices are safe, effective and consistently manufactured.
Two standards have long been central to that process: ISO 13485, which addresses quality management systems for medical devices, and ISO 14971, which addresses medical device risk management.
But they are not the only standards medical device companies may encounter.
Depending on the device, additional standards can influence everything from biocompatibility and packaging to sterilization, software and electrical safety. Understanding which standards apply, and considering them early enough in development, can help prevent costly changes as a device moves toward validation and commercial manufacturing.
ISO 13485: Quality Management Systems
ISO 13485:2016 establishes quality management system requirements specifically for the medical device industry.
The standard provides a framework for controlling activities throughout the device lifecycle, including design and development, supplier management, production, documentation, validation and corrective actions.
For medical device developers, this becomes increasingly important when moving beyond prototyping.
Building a few successful prototypes demonstrates that a design can work. Commercial manufacturing requires demonstrating that the device can be produced consistently under controlled conditions.
That means controlling materials and suppliers, documenting manufacturing processes, managing changes, maintaining traceability and creating records that demonstrate the product was manufactured according to established requirements.
ISO 13485 has also become even more significant for companies commercializing devices in the United States.
As of February 2, 2026, the FDA’s Quality Management System Regulation (QMSR) is in effect. QMSR incorporates ISO 13485:2016 by reference into FDA’s quality system requirements under 21 CFR Part 820, bringing the U.S. regulatory framework into closer alignment with the internationally recognized standard.
For emerging medical device companies, quality systems therefore should not be something considered only after a device is ready for production.
ISO 14971: Medical Device Risk Management
ISO 14971:2019 provides a framework for identifying hazards associated with medical devices, evaluating the associated risks and implementing appropriate risk controls.
Risk management should occur throughout the device lifecycle, beginning during development and continuing through manufacturing and post-market activities.
This can include risks associated with the device design, materials, manufacturing processes, packaging, sterilization and intended use.
The important connection is that risk management does not exist separately from product development.
A risk identified during development may require a design change. A design requirement may create a manufacturing control. A manufacturing risk may require inspection, testing or process validation.
Addressing those relationships early is generally much easier than discovering them after a design has been finalized.
Other Standards That May Apply
ISO 13485 and ISO 14971 provide an important foundation, but the standards applicable to a particular medical device depend on the product itself.
Some commonly encountered standards include:
| Standard | Area |
|---|---|
| ISO 10993 series | Biological evaluation and biocompatibility |
| ISO 11607-1 & -2 | Packaging for terminally sterilized medical devices |
| ISO 11135 | Ethylene oxide sterilization |
| ISO 11137 series | Radiation sterilization |
| IEC 62304 | Medical device software lifecycle processes |
| IEC 62366-1 | Usability engineering |
| IEC 60601 series | Safety and essential performance of medical electrical equipment |
Not every standard applies to every medical device.
A sterile, single-use catheter will have very different considerations from a non-sterile instrument or an electrically powered diagnostic system.
For example, a patient-contacting device may require biological evaluation under the ISO 10993 series. A terminally sterilized device may need packaging developed and validated in accordance with ISO 11607. The sterilization method itself can introduce additional applicable standards.
The key is identifying those requirements early enough that they can influence development rather than becoming obstacles immediately before commercialization.
Why Standards Matter Before Manufacturing
One of the biggest challenges in medical device commercialization is the transition from “the device works” to “the device can be manufactured repeatedly under controlled conditions.”
Those are not the same milestone.
A prototype may use materials, processes or assembly methods that work perfectly well for a handful of devices but are difficult to control at production scale.
As development progresses, companies need to consider questions such as:
- Can the device be assembled consistently?
- Are critical manufacturing processes controlled?
- Can specifications be reliably inspected and tested?
- Are production materials and suppliers appropriately controlled?
- Does the packaging protect the device through sterilization, distribution and shelf life?
- Will sterilization affect materials, adhesives, coatings or device performance?
These questions connect quality, risk management and manufacturing.
They are also why decisions surrounding ISO standards should not be postponed until regulatory submission or production transfer.
Building Toward Commercialization
ISO standards provide more than a regulatory checklist. They create frameworks for developing medical devices in a controlled and repeatable way.
For emerging medical device companies, the goal is not to apply every available standard. It is to determine which requirements apply to the device and when they need to be addressed.
Doing that early can reduce redesigns, validation delays and manufacturing problems later.
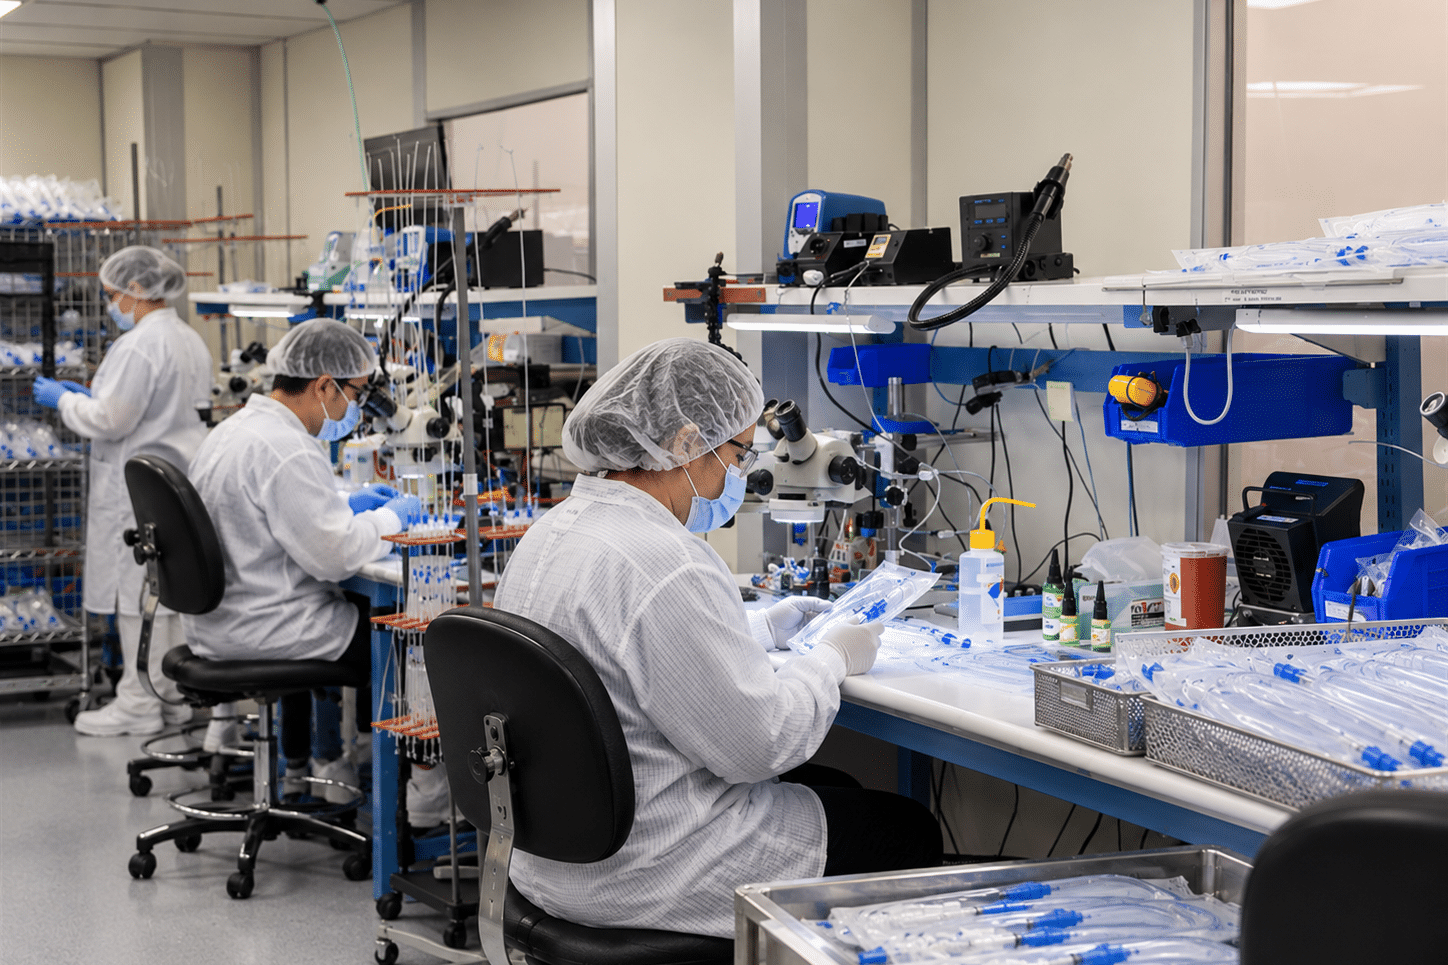
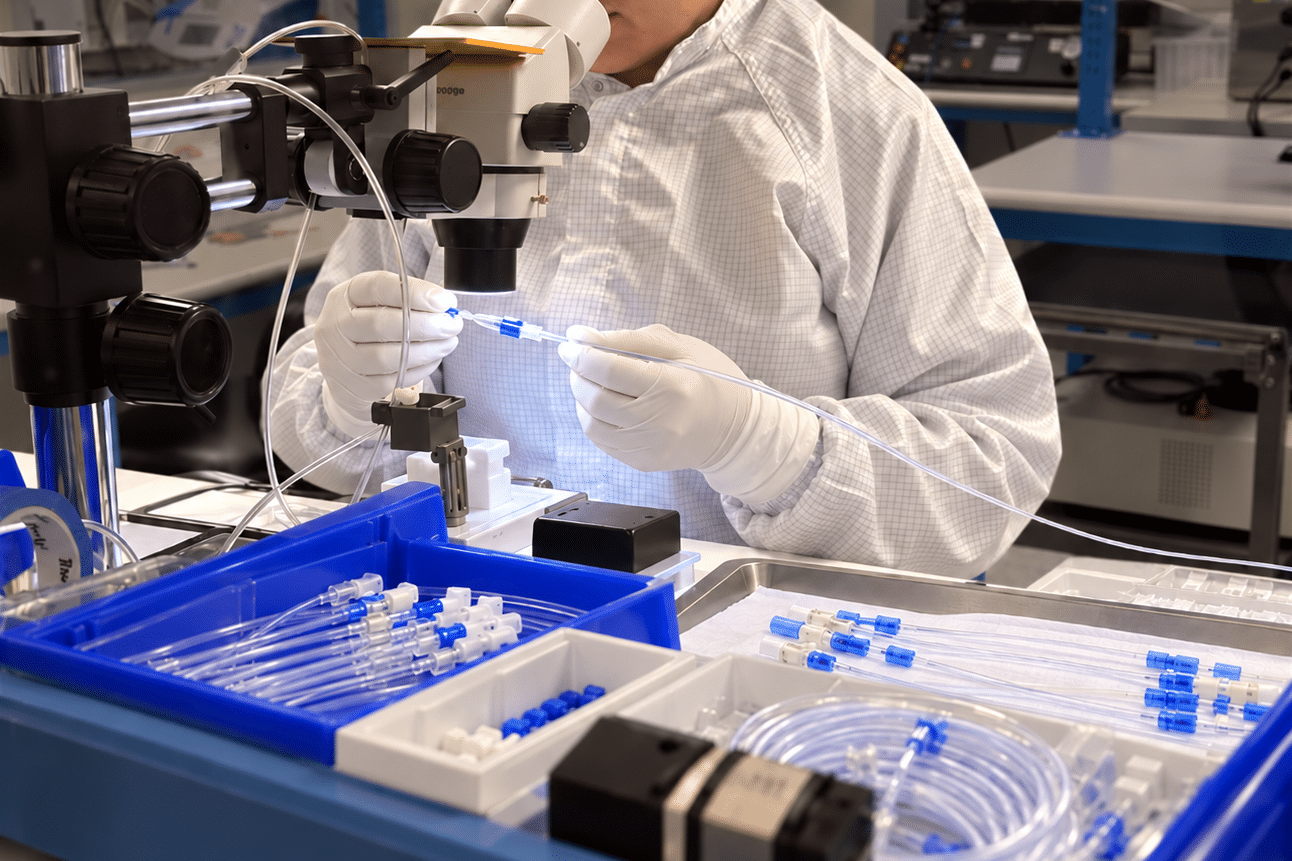
At Pathway Medtech, we help medical device companies make that transition from development into validation and controlled manufacturing. Our capabilities span product development, quality and regulatory support, design verification builds, cleanroom manufacturing, medical device packaging and sterilization coordination.
The earlier development and manufacturing requirements are considered together, the easier it becomes to build a device that is not only ready for regulatory review, but ready to manufacture.
References
- International Organization for Standardization (ISO). ISO 13485:2016 — Medical devices — Quality management systems — Requirements for regulatory purposes. ISO 13485:2016 remains the current edition and was reviewed and confirmed in 2025.
- U.S. Food and Drug Administration (FDA). Quality Management System Regulation (QMSR). Effective February 2, 2026, the QMSR amended 21 CFR Part 820 and incorporated ISO 13485:2016 by reference into FDA’s medical device quality system requirements.
- International Organization for Standardization (ISO). ISO 14971:2019 — Medical devices — Application of risk management to medical devices.
- International Organization for Standardization (ISO). ISO 10993-1:2025 — Biological evaluation of medical devices — Part 1: Requirements and general principles for the evaluation of biological safety within a risk management process.
- International Organization for Standardization (ISO). ISO 11607-1:2019 — Packaging for terminally sterilized medical devices — Part 1: Requirements for materials, sterile barrier systems and packaging systems.
- International Organization for Standardization (ISO). ISO 11607-2:2019 — Packaging for terminally sterilized medical devices — Part 2: Validation requirements for forming, sealing and assembly processes.
- International Organization for Standardization (ISO). ISO 11135:2014 — Sterilization of health-care products — Ethylene oxide — Requirements for the development, validation and routine control of a sterilization process for medical devices.
- International Organization for Standardization (ISO). ISO 11137-1:2025 — Sterilization of health care products — Radiation — Part 1: Requirements for the development, validation and routine control of a sterilization process for medical devices.
- International Electrotechnical Commission (IEC). IEC 62304 — Medical device software — Software life cycle processes.
- International Electrotechnical Commission (IEC). IEC 62366-1 — Medical devices — Part 1: Application of usability engineering to medical devices.
- International Electrotechnical Commission (IEC). IEC 60601 series — Medical electrical equipment.